Study Overview
The study focuses on a unique case involving a young girl from Sudan who presented with a combination of childhood-onset parkinsonism, dystonia, and severe muscle wasting. These symptoms are indicative of significant neurological and muscular dysfunction. As part of the case investigation, the researchers aimed to identify the underlying genetic causes of the girl’s condition. Their findings revealed a novel homozygous variant in the DnaJ heat shock protein family, specifically the heat shock protein 40 member C6. This frameshift mutation is particularly noteworthy as it hasn’t been previously documented, suggesting a possible link to the girl’s clinical presentation.
Additionally, the study adds another layer of complexity by reporting an incidental finding: a pathogenic variant in the breast cancer 1 gene (BRCA1). Such findings are often encountered in genetic testing, raising important considerations regarding the implications for the patient’s health and the potential for familial risk assessment. The emphasis on the DnaJ protein family is critical, as these proteins play a significant role in protein folding and cellular stress responses, which could shed light on the mechanisms of the neurological symptoms observed in the patient.
This research underscores the importance of comprehensive genetic screening in understanding rare congenital disorders, especially in populations where specific genetic variants might be prevalent. Moreover, it highlights the potential for discovering incidental findings that can influence future treatment strategies and screening recommendations for related health concerns, particularly in affected families. The integration of genetic data with clinical observations provides a more detailed understanding of the interplay between genetic mutations and phenotypic expression, which is essential for advancing personalized medicine approaches.
Case Presentation
The patient is a 10-year-old girl from Sudan who presented with a complex array of symptoms that became apparent in early childhood. Over a period of time, she developed signs of parkinsonism, characterized by bradykinesia, rigidity, and a significant postural instability. Alongside these neurological symptoms, she exhibited dystonia, manifesting as involuntary muscle contractions leading to abnormal postures. The severity of her condition was further exacerbated by significant muscle wasting, indicating a profound impact on her muscular function and overall physical health.
The girl’s developmental milestones were notably delayed, aligning with her motor difficulties. Initially, her motor function appeared normal, but the onset of symptoms around the age of 3 raised concerns among her family. These symptoms prompted them to seek medical assistance. Upon examination, healthcare providers noted distinct features, including a tremor in her hands and difficulties in maintaining balance, which are typically associated with parkinsonian disorders. Furthermore, her muscle strength was evaluated, revealing considerable weakness that affected her ability to perform routine activities, contributing to her diminished quality of life.
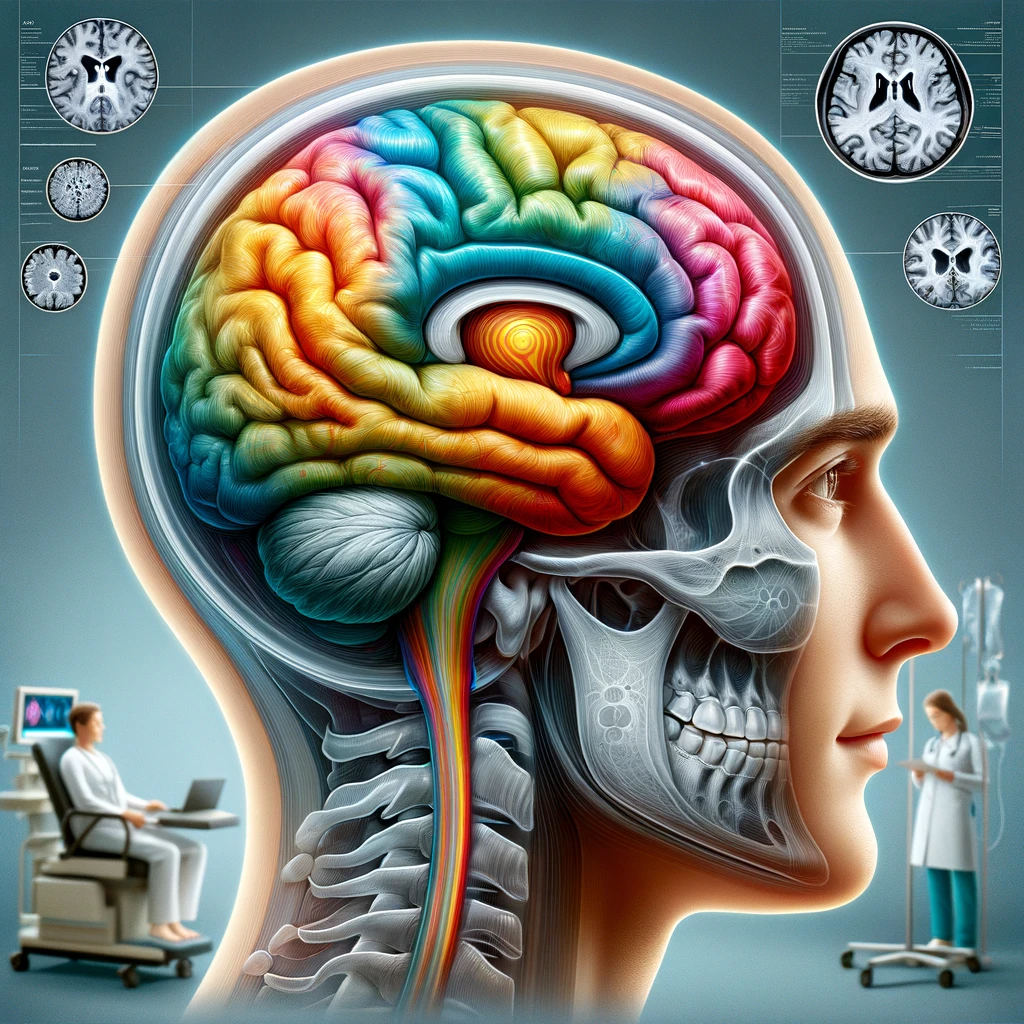
A thorough clinical assessment included neurological evaluations and imaging studies. An MRI of the brain showed subtle changes, although these findings were not specific enough to correlate definitively with a particular diagnosis. The clinical presentation, coupled with her family history, which revealed no prior instances of neurological disorders, warranted a comprehensive genetic evaluation. Genetic counseling was provided to the family, which highlighted the potential implications of hereditary conditions, particularly in light of the girl’s symptoms.
The importance of recognizing incidental findings in genetic screening was illustrated when genetic tests revealed a pathogenic variant in the BRCA1 gene, typically associated with breast and ovarian cancer susceptibility. Although this finding was not directly related to her neurological symptoms, it emphasized the necessity of considering a broad spectrum of health implications for the patient and her family, particularly concerning future cancer risks.
This case underlines the critical need for a multidisciplinary approach in managing complex clinical presentations where genetic factors may play a decisive role. The interplay of the identified genetic variant within the DnaJ family and its relevance to the clinical phenotype provides a compelling example of how specific genetic aberrations can manifest as severe neurological and muscular conditions in children. The discovery of concurrent genetic abnormalities illustrates the complexity of diagnosing and treating multifaceted disorders, reinforcing the value of integrated genetic testing in pediatric medicine.
Genetic Analysis
The genetic analysis of the patient revealed a novel homozygous variant in the DnaJ heat shock protein family, specifically the heat shock protein 40 member C6 (DNAJC6). This particular frameshift mutation presents significant implications, as it alters the genetic coding sequence that is crucial for the proper functioning of the DnaJ proteins. These proteins are integral to cellular processes, particularly in response to stress and the maintenance of protein homeostasis. A dysfunctional DnaJ protein can lead to impaired neuronal health, which may explain the patient’s complex neurological manifestations, including parkinsonism and dystonia.
Homozygosity of the variant suggests that both alleles of the gene are affected, providing a clear linkage to the severe phenotypic expression seen in the patient. This inherited condition implicates not only a genetic predisposition but also highlights the potential role of consanguinity or genetically unique backgrounds within the Sudanese population, where specific gene variants may be more prevalent. The identification of this frameshift mutation contributes to the expanding database of known variants associated with childhood parkinsonism and related disorders, underscoring the need for further studies to investigate the variant’s functional impact.
In addition to the DnaJ variant, incidental findings revealed the presence of a pathogenic variant in the BRCA1 gene. While BRCA1 mutations are predominantly recognized for their strong associations with breast and ovarian cancer, the discovery in this young patient introduces a new dimension of considerations for genetic counseling. The presence of such variants necessitates comprehensive discussions about immediate health implications as well as long-term surveillance and preventive strategies for potential malignancies, not only for the patient but also for family members who may be at risk.
The process of genetic analysis involved whole exome sequencing, a powerful tool that enables the characterization of a wide range of genetic variants across the coding regions of the genome. This technique proved essential in identifying both the causal variant in DNAJC6 and the incidental BRCA1 finding, demonstrating its utility in the diagnostic evaluation of rare diseases. Through this approach, clinicians can obtain a more complete picture of an individual’s genetic landscape, facilitating earlier diagnosis and more tailored therapeutic strategies.
Furthermore, the implications of these genetic findings extend beyond the individual level. They raise important questions regarding family screening and the inheritance patterns of identified variants. The results of genetic testing can influence diagnostic pathways for siblings and extended family members, who may also carry relevant genetic predispositions. The ethical considerations of disclosing incidental findings need to be navigated carefully, ensuring that families have a clear understanding of the potential risks and benefits of such information.
Overall, the genetic analysis highlights the profound role of genetic mutations in modulating clinical presentation, especially in complex cases where multiple conditions may coexist. The intersection of neurological and oncological concerns emphasizes the importance of a multidisciplinary approach in managing pediatric patients with unique genetic profiles and lays the groundwork for future explorations into the mechanisms by which specific mutations contribute to diverse health outcomes.
Discussion and Implications
The case of the Sudanese girl with childhood-onset parkinsonism, dystonia, and muscle wasting highlights the crucial role of genetic factors in the manifestation of complex neurological disorders. The identification of a novel homozygous variant in the DnaJ heat shock protein family underscores the potential for specific genetic mutations to produce severe clinical phenotypes, particularly in pediatric populations. DnaJ proteins are pivotal in regulating cellular responses to stress and maintaining protein stability; therefore, an alteration in their function can lead to significant neurological consequences. The frameshift mutation observed in DNAJC6 suggests that impaired protein function could disrupt normal neuronal health, contributing directly to the symptoms exhibited by the patient.
Additionally, the awareness that the DnaJ variant is rare yet potentially implicated in similar cases emphasizes the value of genetic screening. This could pave the way for identifying at-risk individuals who share similar genetic backgrounds, particularly in populations like those in Sudan, where consanguinity may be more prevalent. Genetic counseling and testing for family members, especially siblings, could be instrumental in early detection and management strategies for hereditary conditions.
The incidental finding of a pathogenic BRCA1 variant adds complexity and depth to the case. While this variant primarily relates to breast and ovarian cancer susceptibility, its discovery prompts vital discussions regarding the broader implications for the patient’s health. Understanding the risk of malignancy is essential not just for the patient but also for family members who might be carriers of the same mutation. This case illustrates the multifaceted nature of genetic testing, where incidental findings can lead to critical preventive health measures.
Moreover, the implications of these findings extend into the realm of personalized medicine. By combining genetic knowledge with clinical presentation, healthcare providers can develop tailored treatment approaches that account for both the primary condition and related health risks. This integrative viewpoint is essential in pediatrics, where the consequences of genetic disorders may evolve as children grow, necessitating ongoing evaluations and adjustments to care plans.
Furthermore, the ethical considerations surrounding incidental findings in genetic testing cannot be overstated. Healthcare professionals must navigate these complexities, ensuring families understand the implications of discovering such variants and providing them with the necessary support and resources to address potential health issues linked to genetic predispositions.
In conclusion, this case exemplifies the importance of a holistic approach to diagnosis and management in rare pediatric neurological disorders. Integrating genetic insights with clinical practice can significantly enhance patient care while providing a framework for future research into the genetic underpinnings of similar conditions. The ongoing exploration of how specific mutations contribute to diverse health outcomes will be paramount in advancing our understanding of both rare and complex diseases within pediatric populations.